Draw a short carbon chain, add an oxygen, and turn the structure in space. The immediate pleasure of Avogadro is that chemistry becomes something you can handle. The more consequential feature comes next: that sketch can become a starting geometry for a calculation. Avogadro is a free, open-source molecular editor and visualization tool whose work spans building structures, preparing inputs, and examining results from other chemistry programs.[1]
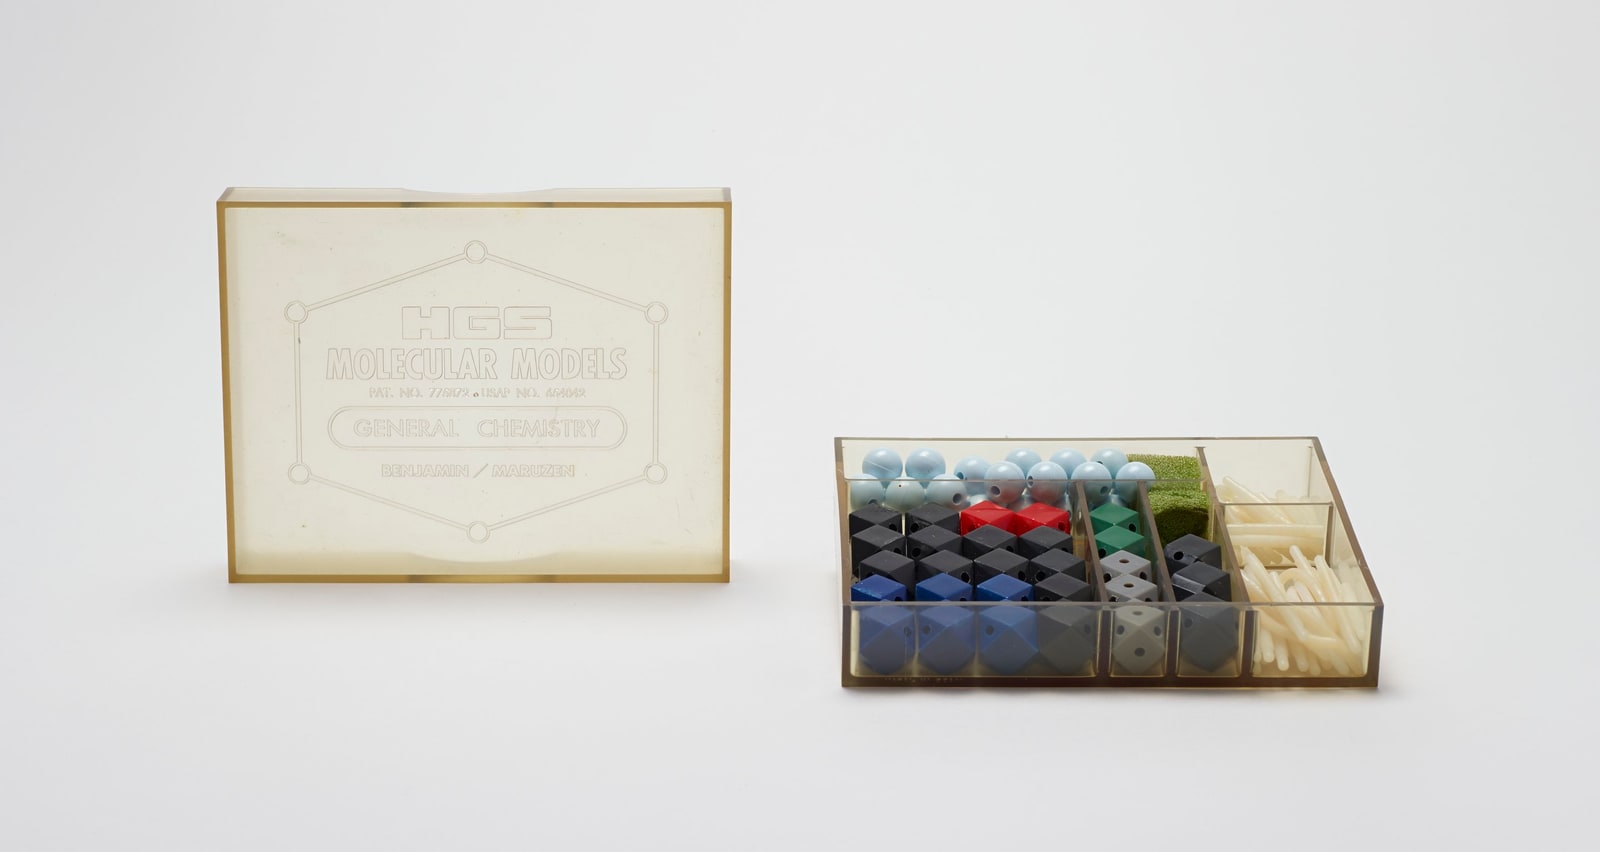
The photograph offers a familiar ancestor of that experience. This HGS model kit, dated 1985–1989 in the Science History Institute's collection, holds colored plastic components in a compartmented container. The museum describes such kits as classroom and laboratory tools for visualizing chemical structures.[2] Avogadro carries the activity onto a computer, where the positions of the pieces can also become data.
There is a timely reason to look again at this established project. Avogadro 2.0.0 arrived on April 1, 2026, with a revised plugin system, an improved ORCA input generator, and updated documentation. Its release notes describe plugins that can add commands, input generators, file formats, and energy models.[3] The interesting proposition is the continuity between making a structure, asking a computational question about it, and returning to inspect the answer.
The drawing already contains assumptions
Avogadro's Draw Tool lets you place an atom with a click and extend a bond by dragging. In version 2.0, its default Automatic bond-order setting uses the distance between atoms to choose single, double, or triple bonds. The Adjust Hydrogens option is enabled by default and adds or removes hydrogen atoms as bonds change.[4]
That saves a great deal of clicking. It also means that a gesture on the screen has chemical consequences. Drag a bond too short and the automatic setting may assign a different bond order than intended. Before optimizing anything, check the element identities, connectivity, and bond orders. The interface makes these choices easy to revise; it still needs the user to know which structure is wanted.
Hydrogens deserve the same attention. The documentation warns that automatic adjustment can interfere with deliberately adding or removing hydrogens to represent an ion or radical. It advises disabling that setting for manual changes.[4] For a beginner, this is a useful first lesson in molecular software: convenience features encode assumptions, and learning where those assumptions live is part of learning the tool.
What happens when the atoms settle
A mouse drawing is an awkward way to specify three-dimensional coordinates. Avogadro's Extensions → Optimize Geometry command adjusts the current structure using a molecular-mechanics force field. Its configuration dialog exposes the chosen force field, convergence criteria, and step limit; selected atoms can also be frozen.[5]
A force field assigns an approximate energy to an arrangement of atoms. Optimization searches for a lower-energy arrangement under that model. The choice of model matters: Avogadro's guide describes MMFF94 as suited to many organic compounds, while UFF covers a wider range of elements. Its simple Lennard-Jones model omits bond-angle, dihedral-angle, and electrostatic terms.[6] A menu containing several methods is therefore offering different descriptions of the system, each with its own scope.
This is where the program becomes more interesting than a digital model kit. The geometry can respond to the selected description of molecular interactions. But the interpretation should stay attached to that description. A tidier structure and a lower displayed energy do not, by themselves, establish the arrangement that would dominate in a particular experiment.
There is another reason to resist stopping at the first tidy result. Flexible molecules can adopt different conformations: arrangements that preserve connectivity while changing their shape. Avogadro documents a separate conformer-search workflow through Open Babel, with systematic, weighted-rotor, and genetic-algorithm options. Its results table compares generated conformers by relative energy and coordinate displacement.[7]
That separation is instructive. Cleaning up one starting arrangement and exploring alternative arrangements answer different questions. If the result depends on the molecule's shape, the practical next step is to examine several candidates and retain the method and settings used to compare them. The lowest value in a particular search remains a result of that search and its energy model.
Give the calculation an explicit question
Avogadro's Input menu provides generators for programs including ORCA, NWChem, CP2K, and Psi4. These generators prepare input for the named programs; the documentation also notes their dependence on a correctly configured Python environment.[8]
The generator interface makes the scientific choices concrete. Avogadro's API documentation identifies fields such as Calculation Type, Theory, Basis, Charge, and Multiplicity.[9] Coordinates say where the atoms start. Those additional choices specify what calculation is being requested and which electronic system it concerns. Generating a file is a handoff to a calculation engine, with further requirements for running and checking that calculation.
This division of work has a long history. In his 2012 Linux Journal review, Joey Bernard presented geometry cleanup as an initial approximation and described exporting inputs to specialized programs before bringing results back for analysis.[10] The article's menus belong to an earlier Avogadro, but its account helps explain the project's continuing appeal. A molecular editor can remain useful across several calculation packages because preparing and inspecting a structure are recurring jobs.
Current Avogadro documentation lists native output parsing for programs including GAMESS, Gaussian, NWChem, and ORCA.[1] That gives the workflow a return journey: the structure that began as a sketch can be examined again in the context of calculated results.
A manageable first experiment
For a student, teacher, or small research group, a sensible introduction is one familiar, small molecule and one intended calculation package. Build the structure, inspect the hydrogens and bonds, optimize with a suitable available model, and examine the generated input before running it. This is a proposed starting exercise, not a claim that a particular chemical method suits every molecule.
For a shared teaching room or laboratory, someone should own the installation and verify the same workflow on the machines people will actually use. Missing Python support can remove expected generators; a generator cannot supply a separately required calculation program simply by naming it.[8] Keep the structure, generated input, program versions, and chosen settings together so the next person can understand the work.
Avogadro's appeal lies in how smoothly a spatial idea becomes something inspectable. You can turn the molecule, revise it, let a model relax it, and pass the coordinates onward. The rewarding habit is to keep asking what each stage has established. That habit turns an attractive molecular picture into a useful beginning.
Sources
- Avogadro, project overview — open-source editor, visualization, and supported calculation-output readers.
- Science History Institute, “HGS General Chemistry Molecular Model Set” — object dated 1985–1989, photograph dated 2016, collection description and image credit.
- Avogadro, “Avogadro 2.0.0,” April 1, 2026 — release notes for plugins, input generation, and documentation.
- Avogadro 2.0 documentation, “Draw Tool” — atom placement, automatic bond order, and hydrogen-adjustment behavior.
- Avogadro 2.0 documentation, “Extensions Menu” — geometry optimization, configuration, and frozen atoms.
- Avogadro 2.0 documentation, “Introduction to Force Fields” — scope of LJ, UFF, and MMFF94 models.
- Avogadro 2.0 documentation, “Finding Conformers of Molecules” — Open Babel searches and comparison of generated conformations.
- Avogadro 2.0 documentation, “Input Menu” — calculation-program interfaces and the Python requirement.
- Avogadro 2.0 API documentation, “Avogadro::MoleQueue::InputGenerator” — calculation type, theory, basis, charge, and multiplicity parameters.
- Joey Bernard, “Basic Chemistry on the GNOME Desktop,” Linux Journal, January 20, 2012 — independent review of the earlier editor-to-calculation workflow; historical context, not current menu instructions.